fastRanges is a multithreaded interval engine for IRanges and GRanges.
It keeps Bioconductor-style overlap semantics and familiar argument grammar
while targeting the workloads that usually dominate runtime in genomics:
large findOverlaps() jobs, repeated query batches against one subject, and
overlap-derived summaries such as counts, joins, and aggregation.
Website: https://cparsania.github.io/fastRanges/
Source: https://github.com/cparsania/fastRanges
if (!requireNamespace("BiocManager", quietly = TRUE)) {
  install.packages("BiocManager")
}
BiocManager::install("fastRanges")
if (!requireNamespace("remotes", quietly = TRUE)) {
  install.packages("remotes")
}
remotes::install_github("cparsania/fastRanges", ref = "main")
library(fastRanges)
library(GenomicRanges)

data("fast_ranges_example", package = "fastRanges")
query <- fast_ranges_example$query
subject <- fast_ranges_example$subject

# One-off overlap call
hits <- fast_find_overlaps(query, subject, threads = 4)

# Repeated-query workflow
subject_index <- fast_build_index(subject)
hits_indexed <- fast_find_overlaps(query, subject_index, threads = 4)

# Derived summaries
counts <- fast_count_overlaps(query, subject_index, threads = 4)
joined <- fast_overlap_join(query, subject, threads = 4)
The package ships a small in-memory example object and matching BED files:
data("fast_ranges_example", package = "fastRanges")
names(fast_ranges_example)

system.file("extdata", "query_peaks.bed", package = "fastRanges")
system.file("extdata", "subject_genes.bed", package = "fastRanges")
fast_find_overlaps(): return overlap pairs as Hits
fast_count_overlaps(): per-query overlap countsfast_overlaps_any(): per-query logical overlap flagfast_build_index(): build a reusable subject indexfast_overlap_join(): overlap join with join = "inner" or "left"
fast_inner_overlap_join(), fast_left_overlap_join()
fast_semi_overlap_join(), fast_anti_overlap_join()
fast_nearest(), fast_distance_to_nearest()
fast_precede(), fast_follow()
fast_count_overlaps_by_group()fast_overlap_aggregate()fast_window_count_overlaps()fast_self_overlaps(), fast_cluster_overlaps()
fast_reduce(), fast_disjoin(), fast_gaps()
fast_range_union(), fast_range_intersect(), fast_range_setdiff()
fast_coverage()fast_tile_coverage()fast_save_index(), fast_load_index(), fast_index_stats()
fast_find_overlaps_iter()fast_iter_has_next(), fast_iter_next()
fast_iter_reset(), fast_iter_collect()
fastRanges is designed to stay close to Bioconductor overlap semantics for
supported inputs, but it is currently best viewed as a high-throughput engine
for IRanges and GRanges, not as a blanket replacement for every
findOverlaps() input class.
Currently supported:
IRangesGRangesselect = "all", "first", "last", and "arbitrary"
Currently unsupported:
GRangesListUnsupported inputs are rejected explicitly with a clear error.
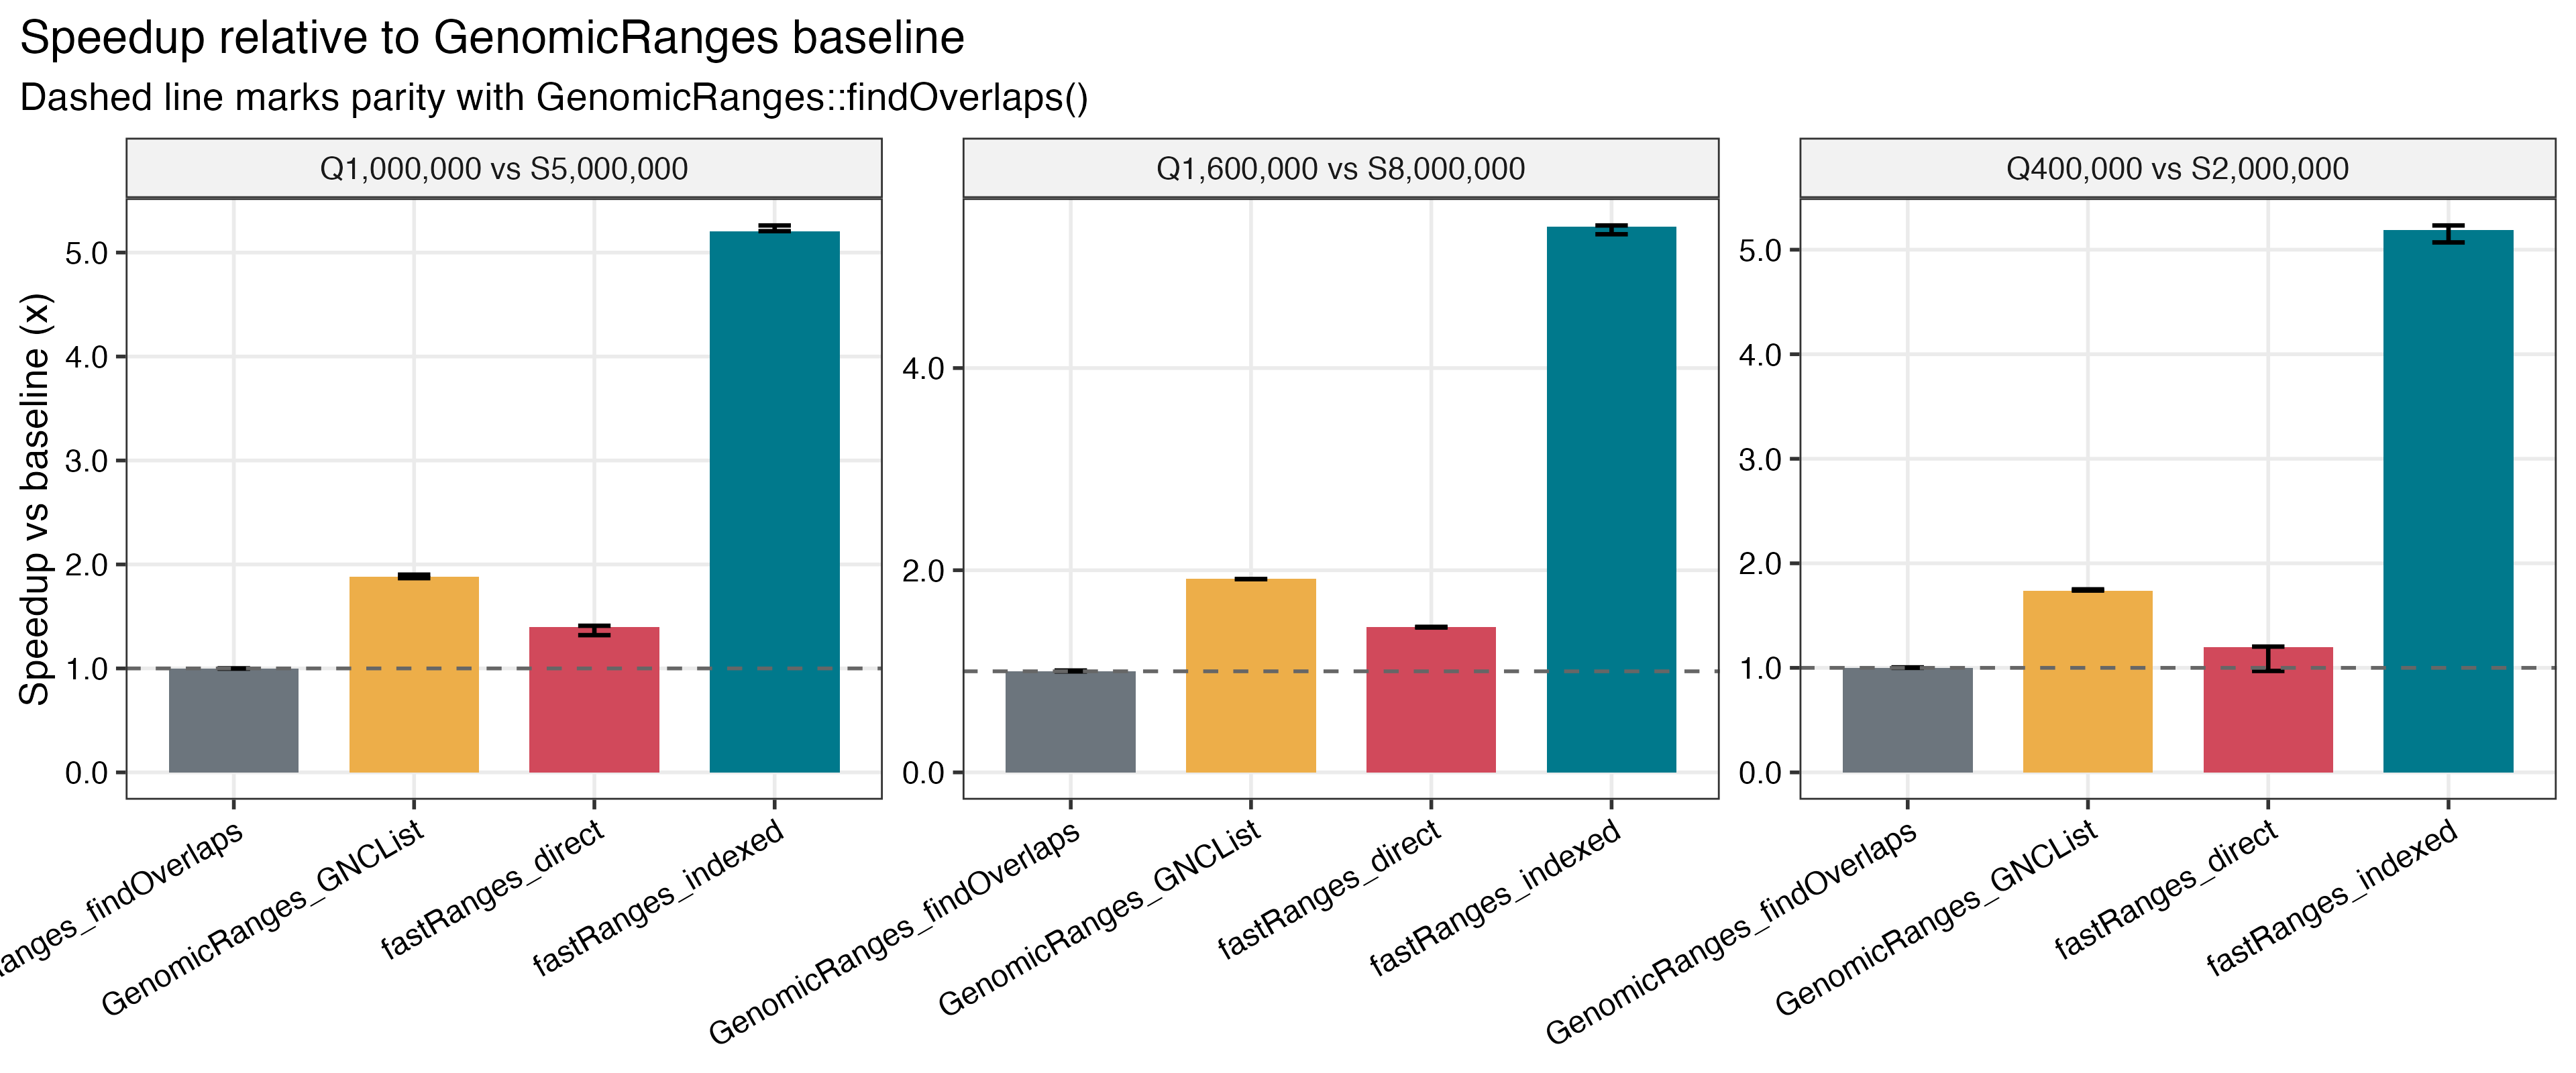
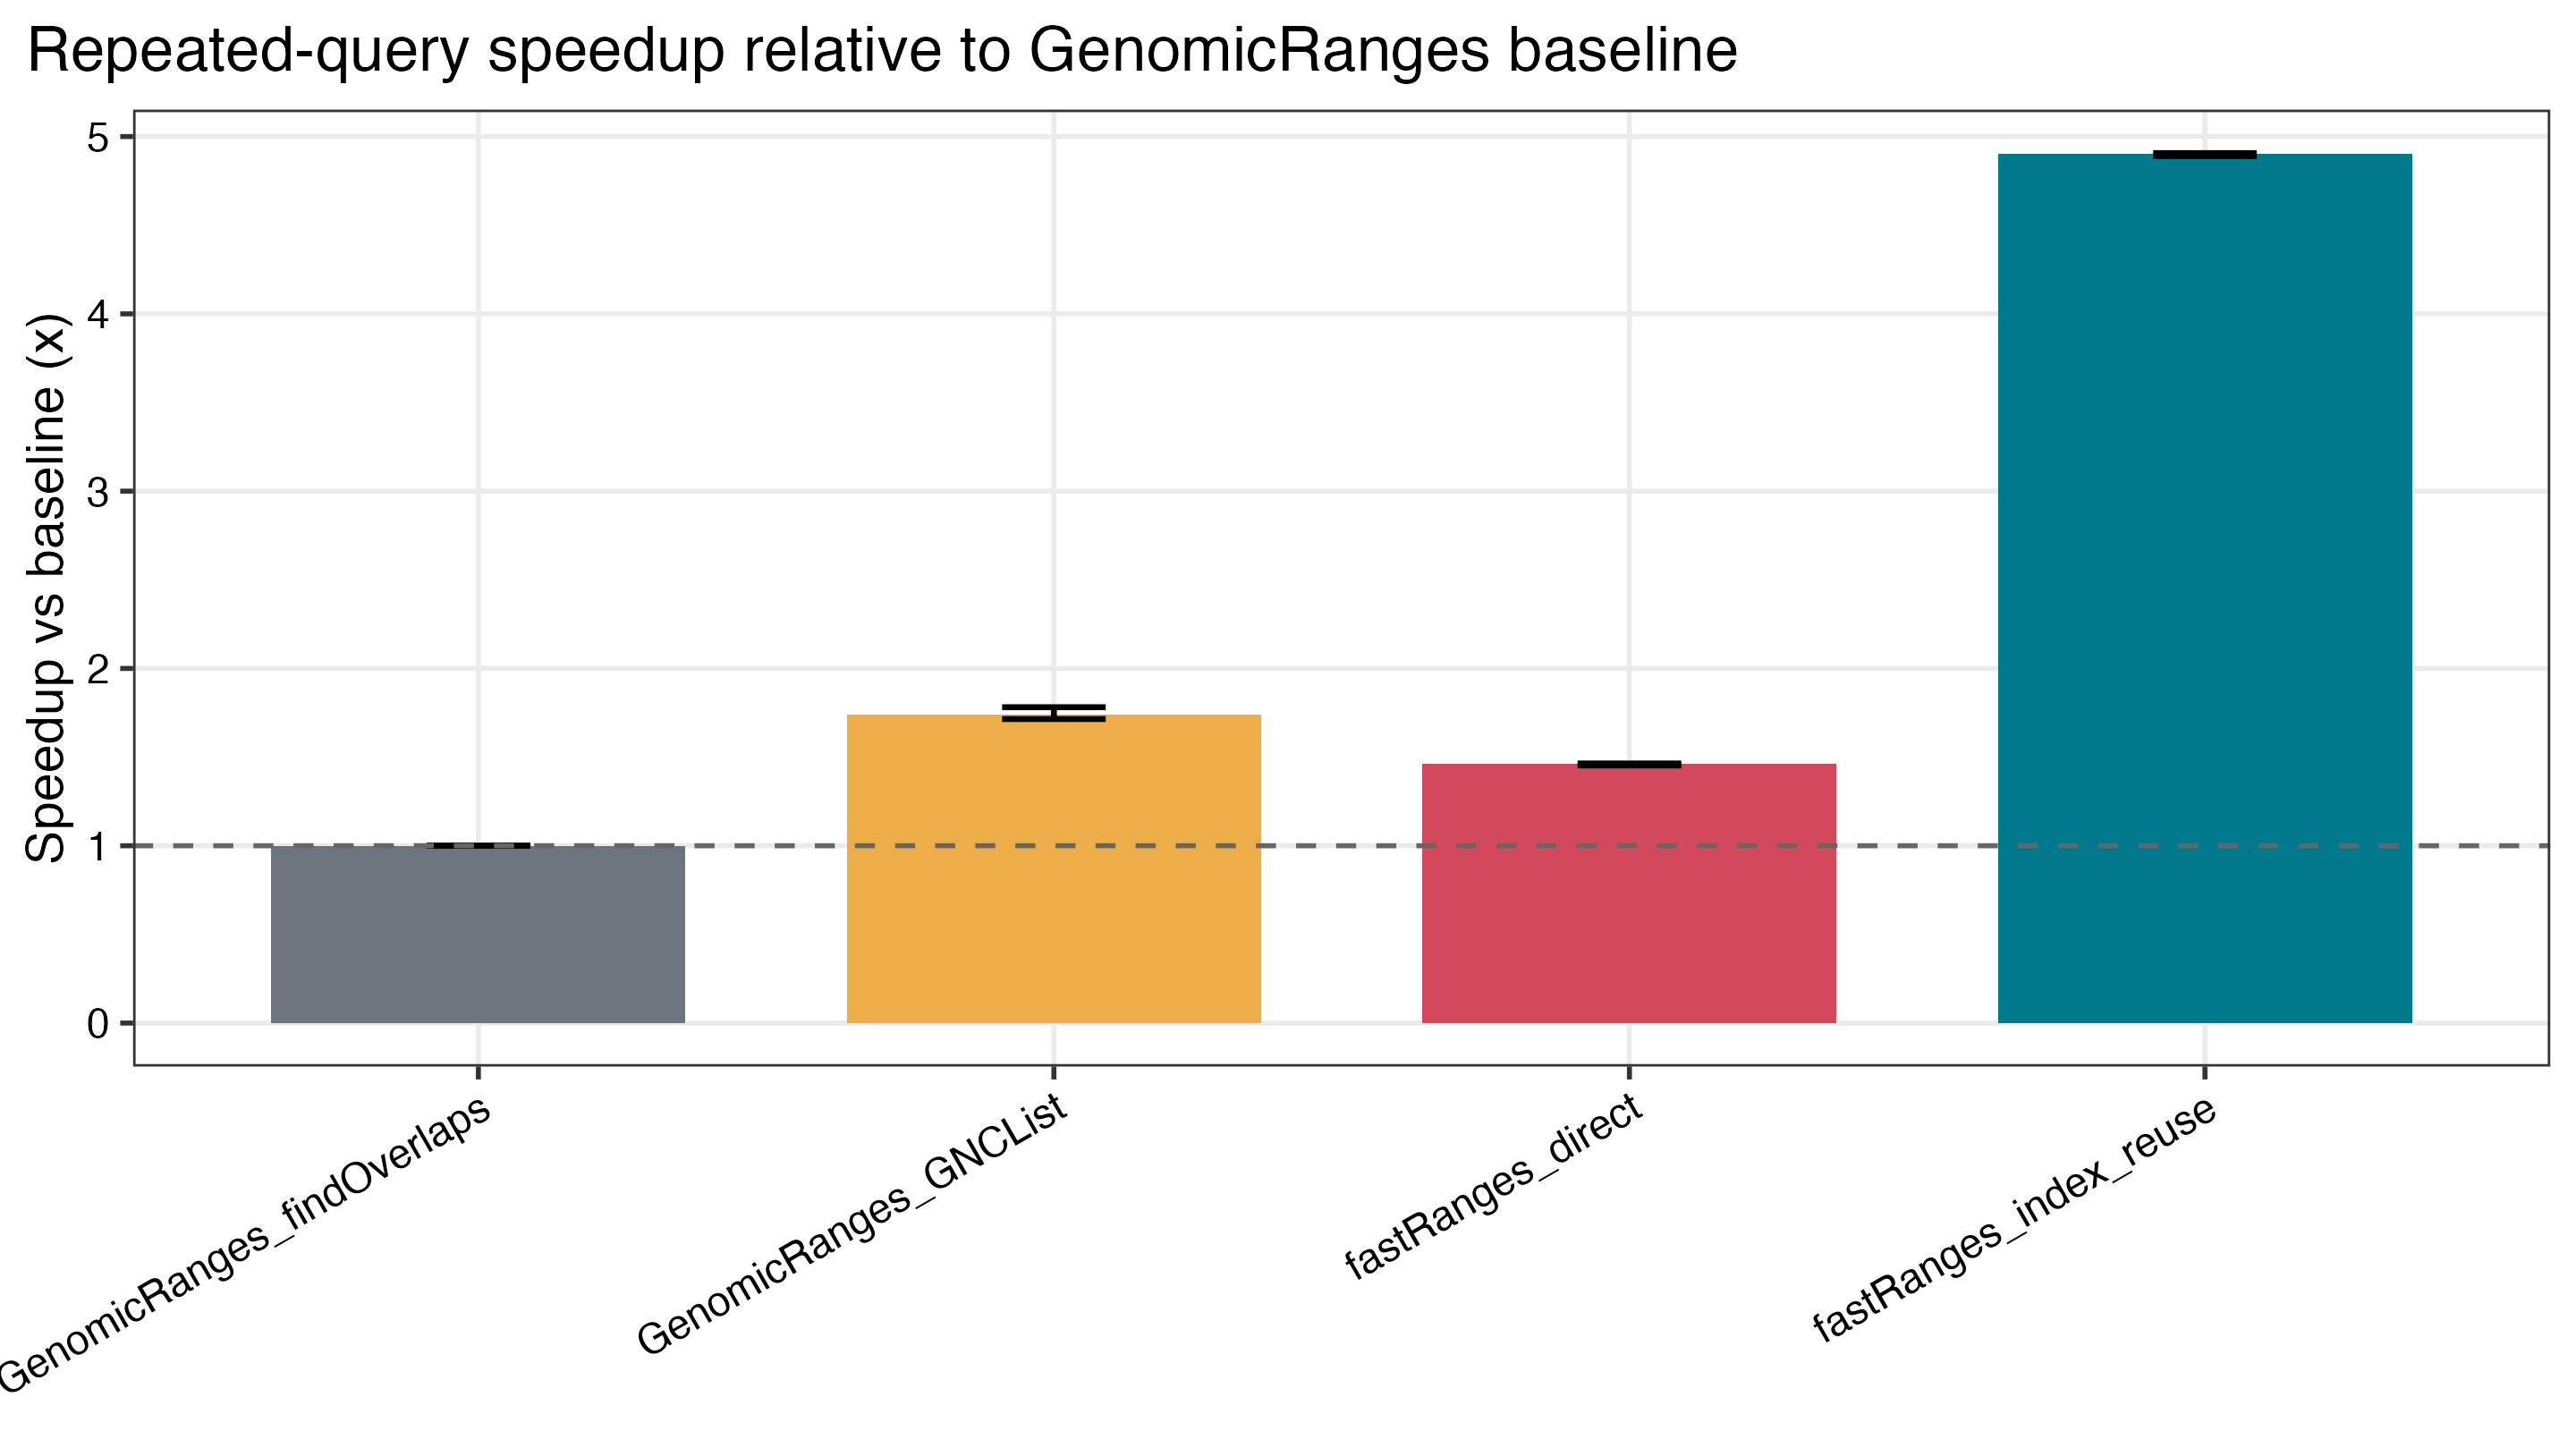
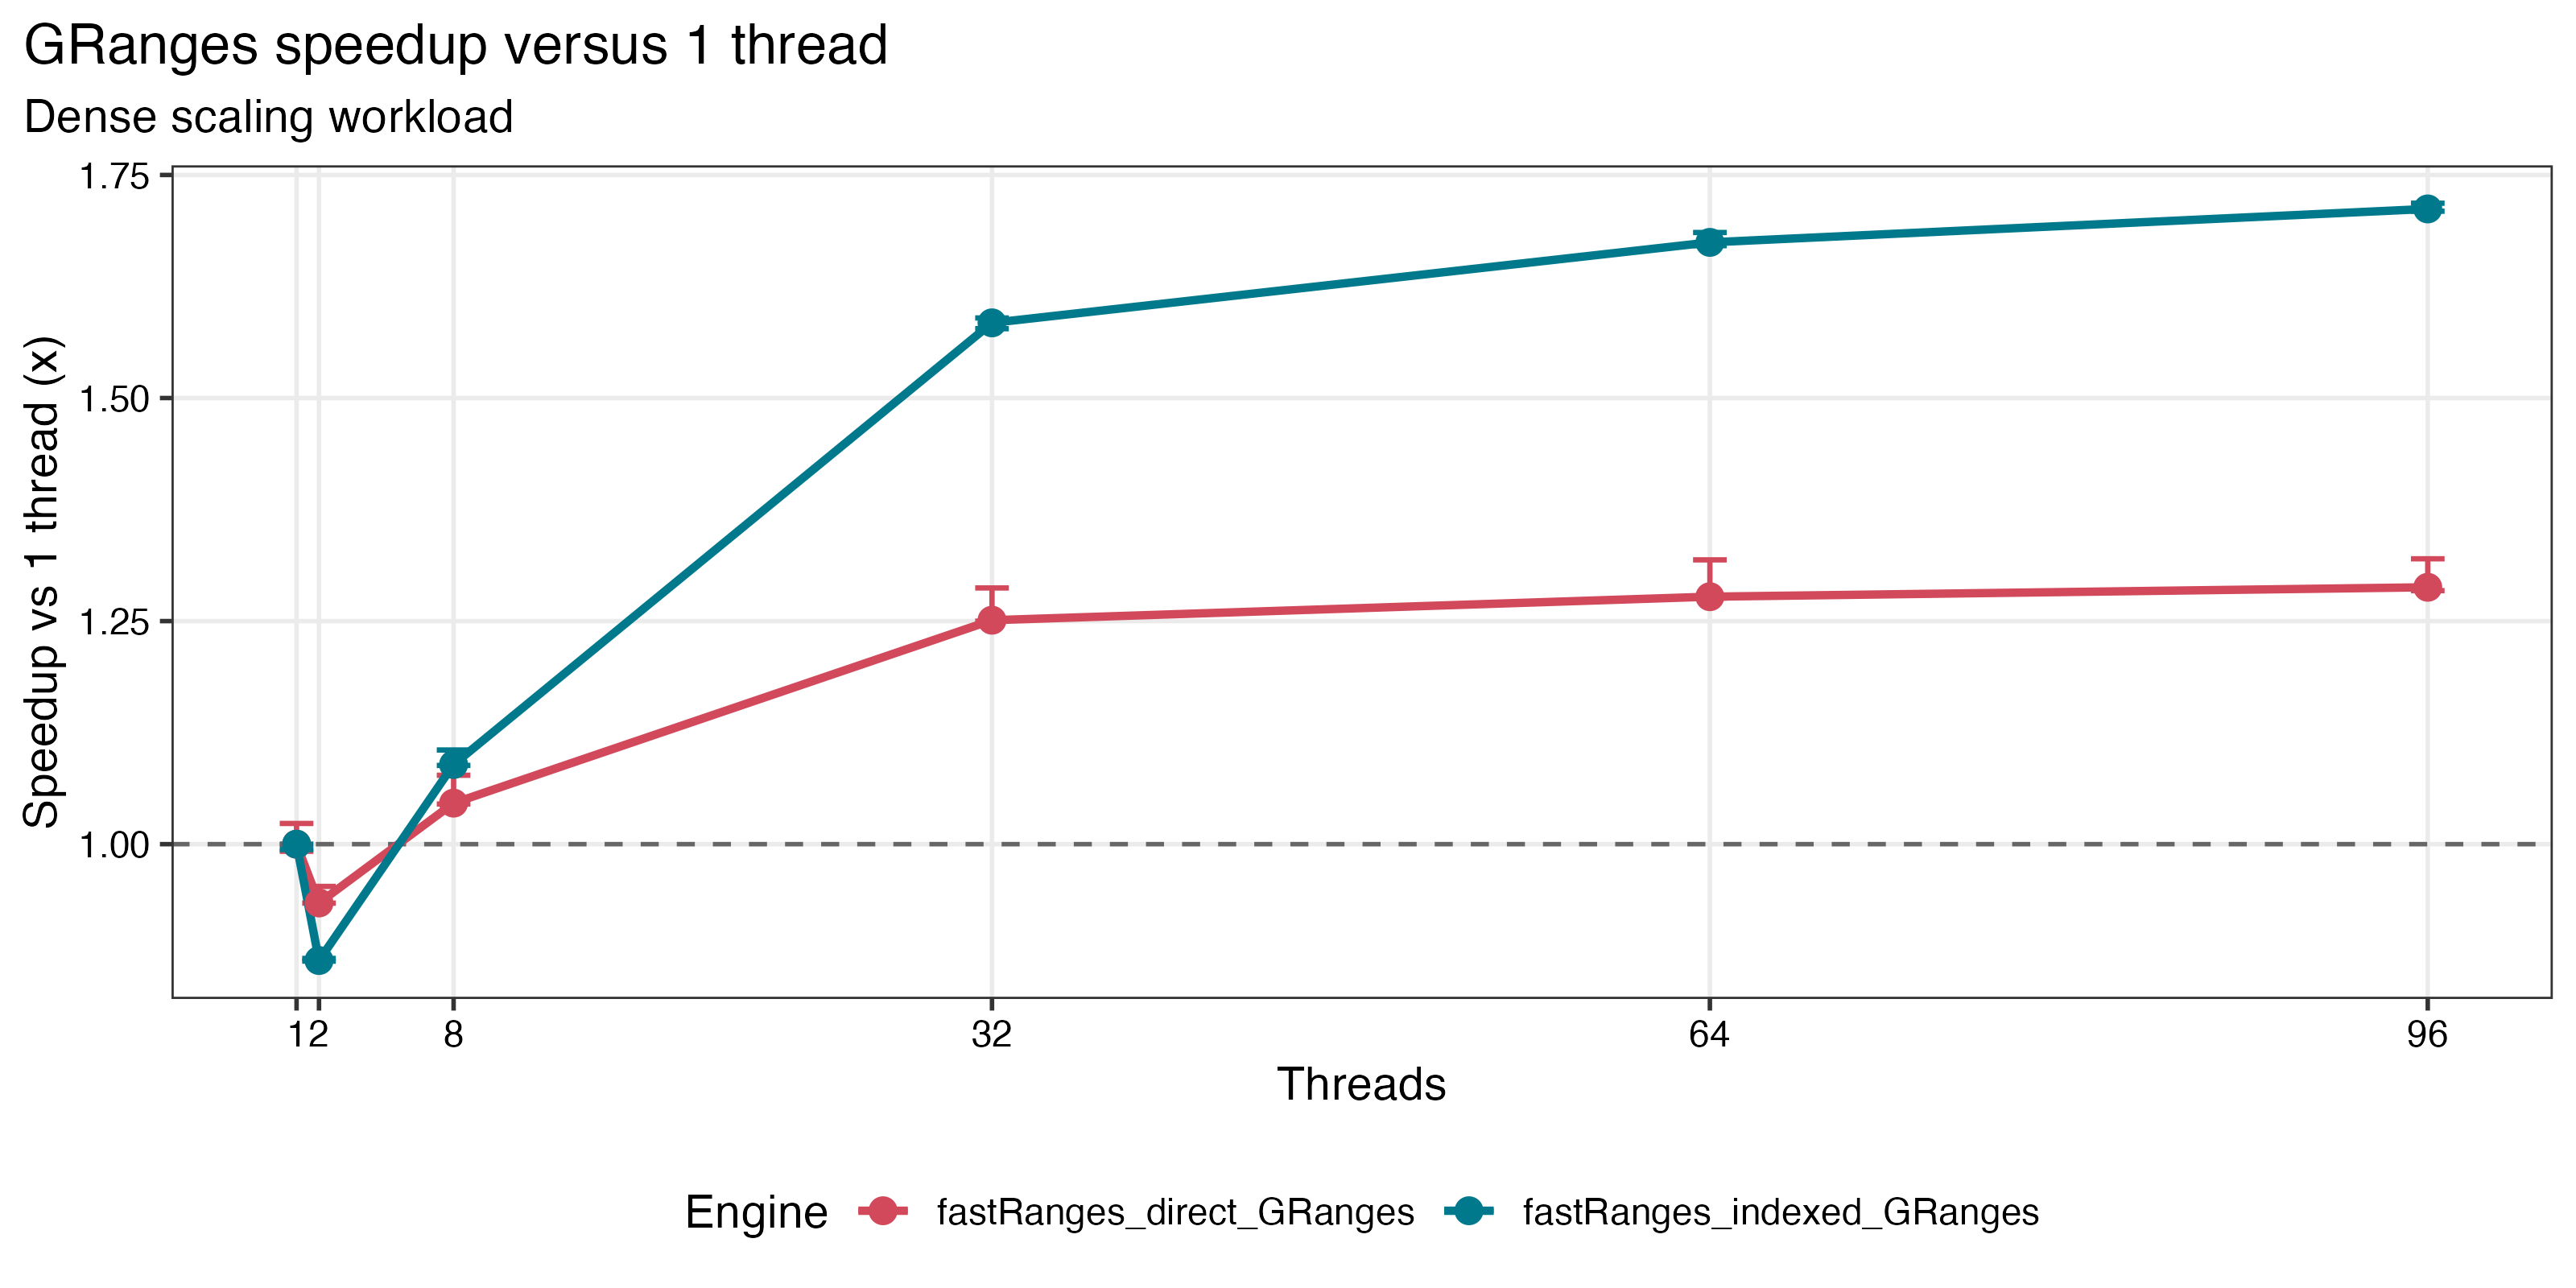
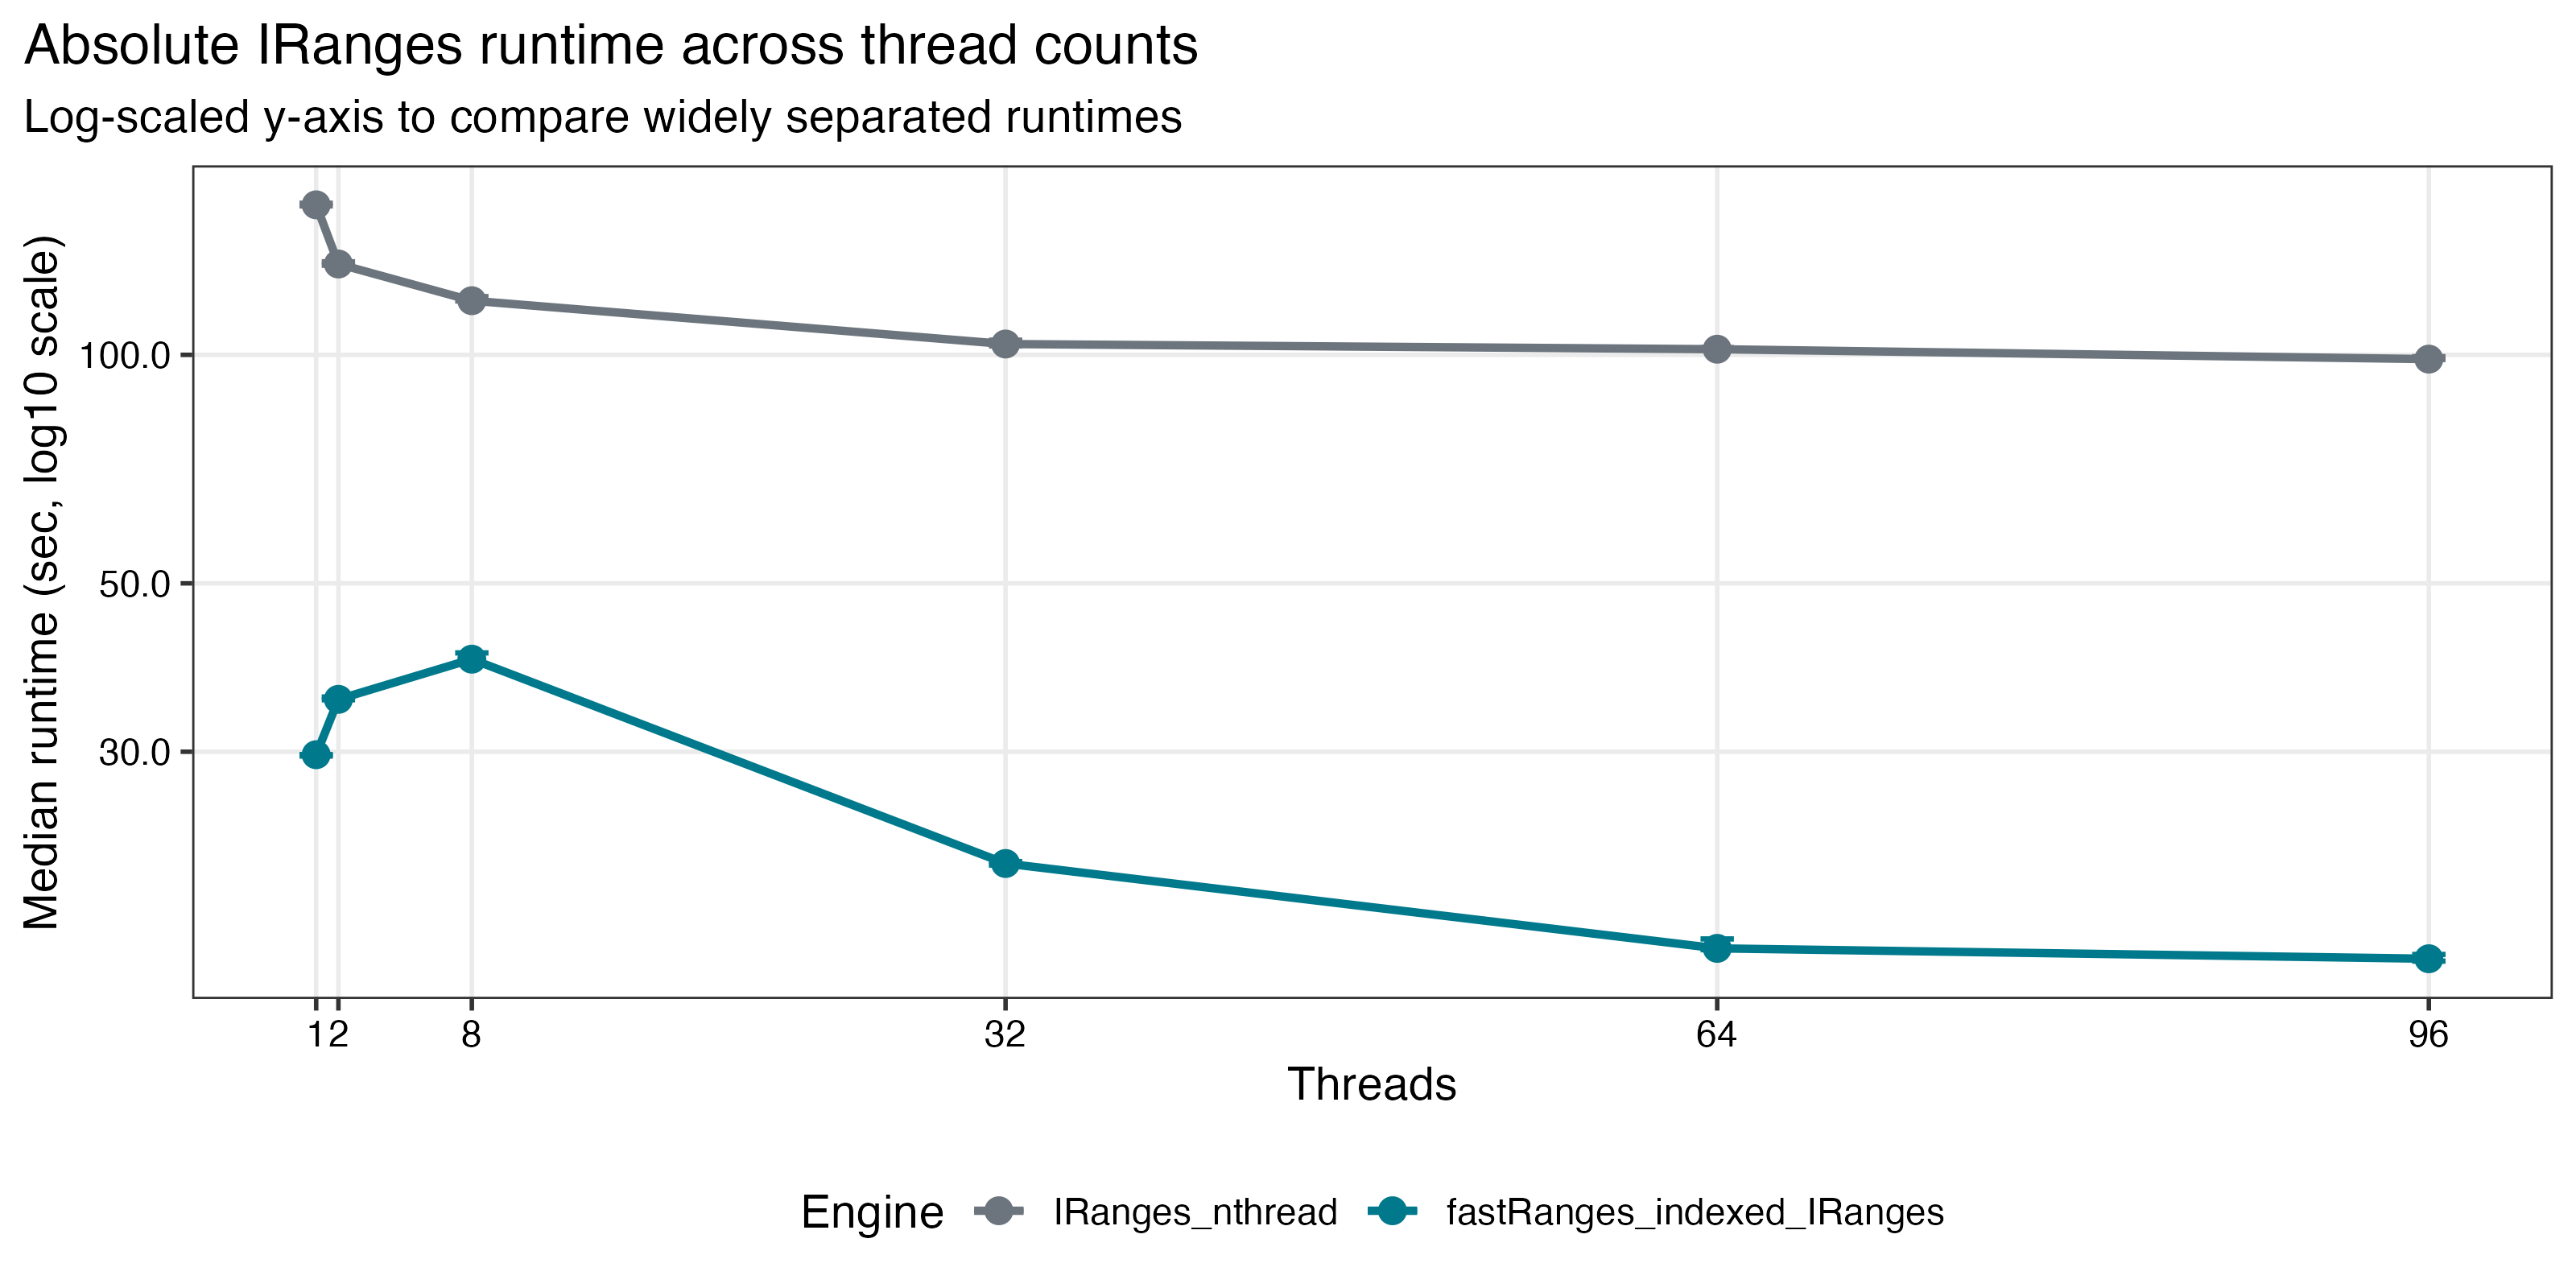
Saved benchmark results on a 96-core Linux server show:
5.19x to 5.40x GRanges speedup for indexed fastRanges versus
GenomicRanges::findOverlaps()
4.90x speedup in repeated-query workloads when the subject index is
reusedGRanges and large IRanges workloads| GRanges speedup vs baseline | Repeated-query speedup |
|---|---|
|
|
| Dense GRanges scaling | IRanges absolute runtime |
|---|---|
 |
 |
Benchmark resources:
fast_build_index(subject) when the same annotation is queried many
times.threads for large workloads on multicore machines.deterministic = TRUE when stable output ordering matters.deterministic = FALSE when maximum multithreaded throughput matters
more than stable hit ordering.